Skip to content
  • Home
  • Recent
Collapse
Brand Logo
CRYSTAL23
Latest v1.0.1
Tutorials Try the Demo Get a License
Tutorials Try the Demo Get a License Instagram

Developer

Private

Developer of the CRYSTAL code!

Posts


  • OPTGEOM fail during ELASTCON
    aerba aerba
    Elasticity, Piezoelectricity, Photoelasticity

    Hi,

    The keyword PRESSURE requires the keyword PREOPTGEOM to pre-optimize the structure at the desired pressure of 15 GPa.

    It is recommended to increase the level of printing in the output with the PRINT keyword to better see what is happening.

    Please, try with the following input:

    ELASTCON
    PRINT
    PREOPTGEOM
    PRESSURE
    15
    END
    END

    and let me know what you get.


  • Einsteinium problem: very small Es DFT radius and discrepancy in the integrated electron number
    aerba aerba
    Bug Reports

    Dear Marco,

    Thank you for your very detailed analysis. We had a look in the code and can indeed confirm that this is a bug.

    For "historical reasons", atomic radii for DFT integration in CRYSTAL are tabulated up to atomic number 98 (Cf). Einsteinium (99) is the first excluded element. Unfortunately, its radius can not be user-defined via the keyword RADIUS because in CRYSTAL atomic number 99 meant "I'm done!" (as you may have seen in the basis set input block which ends with a string "99 0").

    So I am afraid there is no way to run a meaningful DFT calculation with CRYSTAL23 for Es. We are now working at a bug-fix that we will include in future versions of the code.

    Sorry not to have better news this time around, and
    Thanks for reporting this issue!


  • convergence issue with an insulator
    aerba aerba
    Single-Point Calculations

    Can you try with:

    NODIIS
    LEVSHIFT
    10 1
    FMIXING
    80
    

    and let me know if it improves things?


  • ERROR **** INPBAS **** NO BASIS FUNCTION FOR ATOMIC NUMBER 0
    aerba aerba
    Basis Sets

    Hi,

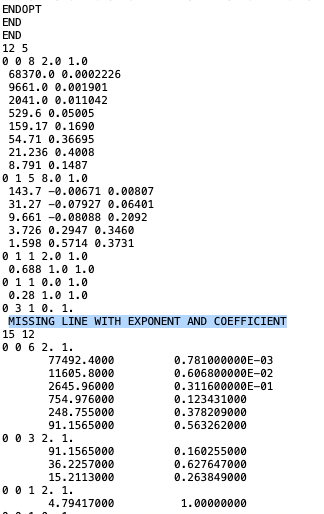
    I could spot the mistake: there is a missing line for the exponent of the d-type shell of Mg:Basis_set_problem.png


  • Presenting CRYSTALLine beta: build, view and edit CRYSTAL structures on your desktop
    dmitoli dmitoli
    CRYSTALLine

    Dear CRYSTAL community,

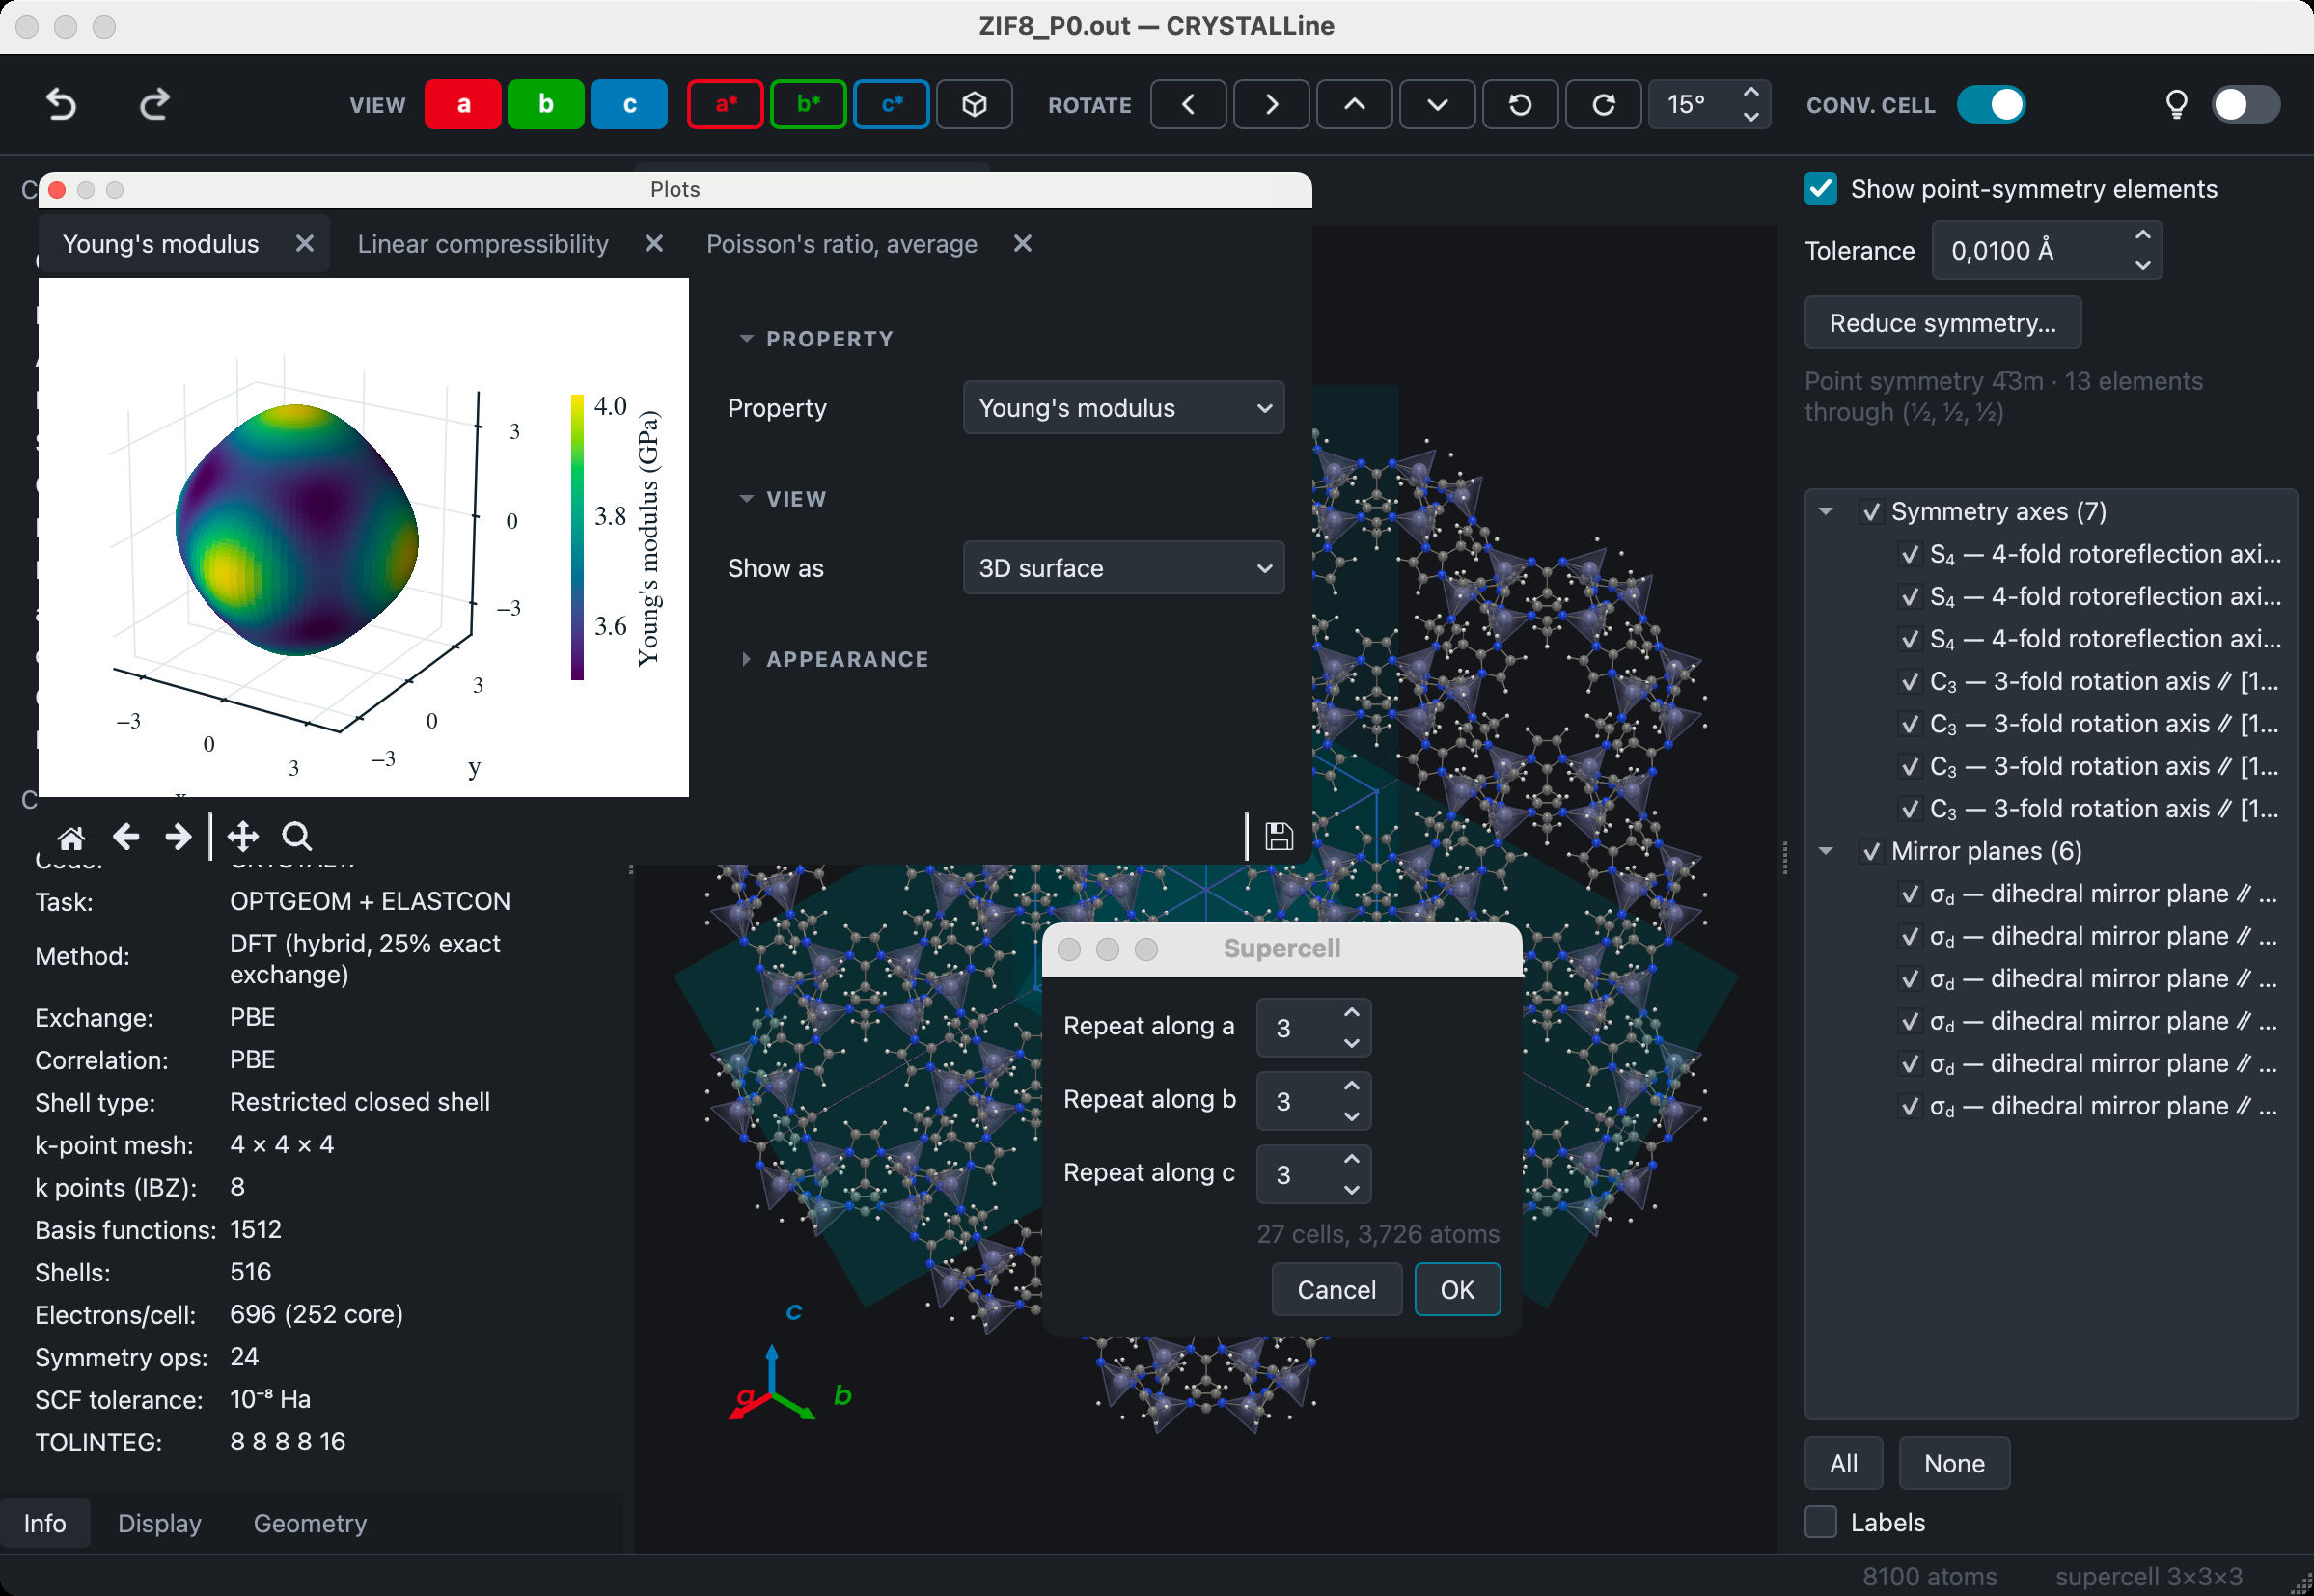
    We would like to introduce the beta release of CRYSTALLine, a new desktop app to display and edit structures for CRYSTAL, to animate phonon modes, and to plot the properties a calculation returns. It's cross-platform: it runs on Linux, macOS and Windows. It is built on CRYSTALClear, and is free software under the GNU GPL v3.

    CRYSTALLine follows in the tradition of MOLDRAW, written by Prof. Piero Ugliengo, which served this community for over twenty years: a program on your own machine for looking at a structure and editing it.

    screen3.png

    What it does

    • Handling of CRYSTAL outputs, .gui/.f34 and CIF files, each in a tab of its own, with crystallographic and calculation data beside it.
    • Automated live detection of symmetry (through spglib).
    • Easy structure editing through drag-and-drop: add, move, delete or change atoms, build supercells, switch between primitive and crystallographic settings.
    • Animation of vibrational modes of a frequency calculation, including out-of-Γ phonons with several exporting options.
    • Input builder for CRYSTAL (.d12) and PROPERTIES (.d3) from the structure on screen.
    • First Brillouin Zone visualiser and k-path selection.
    • Advanced plotting tools (from CRYSTALClear): electronic and phonon band structure, densities of states, IR and Raman spectra (harmonic and anharmonic), elastic surfaces and sections, equations of state, and simulated XRD patterns. Most plots have their options beside them and are redrawn as these are changed.
    • Visualisation of crystalline orbitals, charge density and electrostatic potential over the structure as isosurfaces or as slices along a lattice plane.

    Installing

    It needs Python 3.11 or newer, best in an environment of its own:

    conda create --name crystal python=3.12
    conda activate crystal
    pip install CRYSTALLine
    crystalline
    

    The documentation has a small MgO example to try it, with a band structure and a phonon dispersion.

    This is a beta

    Accidents may take place. We would be grateful for reports of anything that fails or looks wrong, either in reply to this post or at github.com/crystaldevs/CRYSTALLine/issues. Suggestions for what to add next are more than welcome too.

    Links

    Documentation: crystaldevs.github.io/CRYSTALLine
    Source code: github.com/crystaldevs/CRYSTALLine
    PyPI: pypi.org/project/CRYSTALLine

    Give it a try!


  • ELASTCON problem
    aerba aerba
    Elasticity, Piezoelectricity, Photoelasticity

    A possible "work-around" just came to my mind. As far as I understand, your job successfully completed the first deformation (STRAIN 1) to later fail during the next deformation (STRAIN 3) with the following error:

    ERROR **** SCF **** SG_IRR ALLOCATION ERROR: ALREADY ALLOCATED

    It looks like some arrays are not properly deallocated when moving from one deformation to the next. A possible work-around would be to compute each deformation in an independent calculation. You can use the DEFORM option to specify what deformations should be active. For instance with:

    ELASTCON
    PRINT
    DEFORM
    0 0 1 0 0 0
    END
    

    you would be asking to consider just the third deformation as active. With PRINT you ask for extended printing options. You will find the computed elastic constants for the third row of the elastic tensor printed in the output.

    As for your calculation there are only three symmetry-independent deformations:

    LIST OF ACTIVE DEFORMATIONS:
    1
    3
    4

    you could have all the elastic constants by running three calculations. One with:

    ELASTCON
    PRINT
    DEFORM
    1 0 0 0 0 0
    END
    

    a second one with

    ELASTCON
    PRINT
    DEFORM
    0 0 1 0 0 0
    END
    

    and a third with

    ELASTCON
    PRINT
    DEFORM
    0 0 0 1 0 0
    END
    

    Let me know if this helps.

  • ELASTCON problem
    aerba aerba
    Elasticity, Piezoelectricity, Photoelasticity

    Hi,

    We have run your input files with CRYSTAL23 and the calculation was successful (see attached output file). Are you by any chance running an older version of the program?


  • ELASTCON problem
    aerba aerba
    Elasticity, Piezoelectricity, Photoelasticity

    Let me run some tests


  • Necessary memory estimation
    aerba aerba
    Other Questions

    Hi,

    While there is no option to estimate the necessary memory, there are a couple of advanced printing options to monitor the memory used step-by-step throughout the calculation. Please, refer to the MEMOPRT and MEMOPRT2 keywords in the User's Manual.

    Let me know if this helps,


  • Regarding ".out" file not being updated despite job status "running"
    aerba aerba
    Other Questions

    Hi,

    The input/output files you uploaded look fine. It is difficult for me to understand what happened.

Member List

CrystalSupport CrystalSupport Group Owner
0 Posts 0 Reputation
ldona ldona Group Owner
2 Posts 2 Reputation
Chiara Chiara Group Owner
0 Posts 0 Reputation
Jacques Jacques Group Owner
9 Posts 12 Reputation
bcivalleri bcivalleri Group Owner
1 Posts 0 Reputation
aerba aerba Group Owner
119 Posts 95 Reputation
SilviaCasassa SilviaCasassa Group Owner
6 Posts 4 Reputation
dmitoli dmitoli Group Owner
12 Posts 24 Reputation
GiacomoAmbrogio GiacomoAmbrogio Group Owner
74 Posts 80 Reputation
  • Login

  • Don't have an account? Register

  • Login or register to search.
  • First post
    Last post
0
  • Home
  • Recent