Skip to content
  • 97 Topics
    442 Posts
    aerba
    Hi, The keyword PRESSURE requires the keyword PREOPTGEOM to pre-optimize the structure at the desired pressure of 15 GPa. It is recommended to increase the level of printing in the output with the PRINT keyword to better see what is happening. Please, try with the following input: ELASTCON PRINT PREOPTGEOM PRESSURE 15 END END and let me know what you get.
  • 29 Topics
    101 Posts
    fbodo
    Dear @drmajouri2025, Could you provide us your .d3, .outp, and eventually the .f9 or the related .d12, so that we can try to find the issue?
  • Seek assistance, discuss troubleshooting tips for any technical problem you encounter and report bugs

    19 92
    19 Topics
    92 Posts
    aerba
    Dear Marco, Thank you for your very detailed analysis. We had a look in the code and can indeed confirm that this is a bug. For "historical reasons", atomic radii for DFT integration in CRYSTAL are tabulated up to atomic number 98 (Cf). Einsteinium (99) is the first excluded element. Unfortunately, its radius can not be user-defined via the keyword RADIUS because in CRYSTAL atomic number 99 meant "I'm done!" (as you may have seen in the basis set input block which ends with a string "99 0"). So I am afraid there is no way to run a meaningful DFT calculation with CRYSTAL23 for Es. We are now working at a bug-fix that we will include in future versions of the code. Sorry not to have better news this time around, and Thanks for reporting this issue!
  • Discuss tools and techniques for visualizing simulated data

    6 20
    6 Topics
    20 Posts
    dmitoli
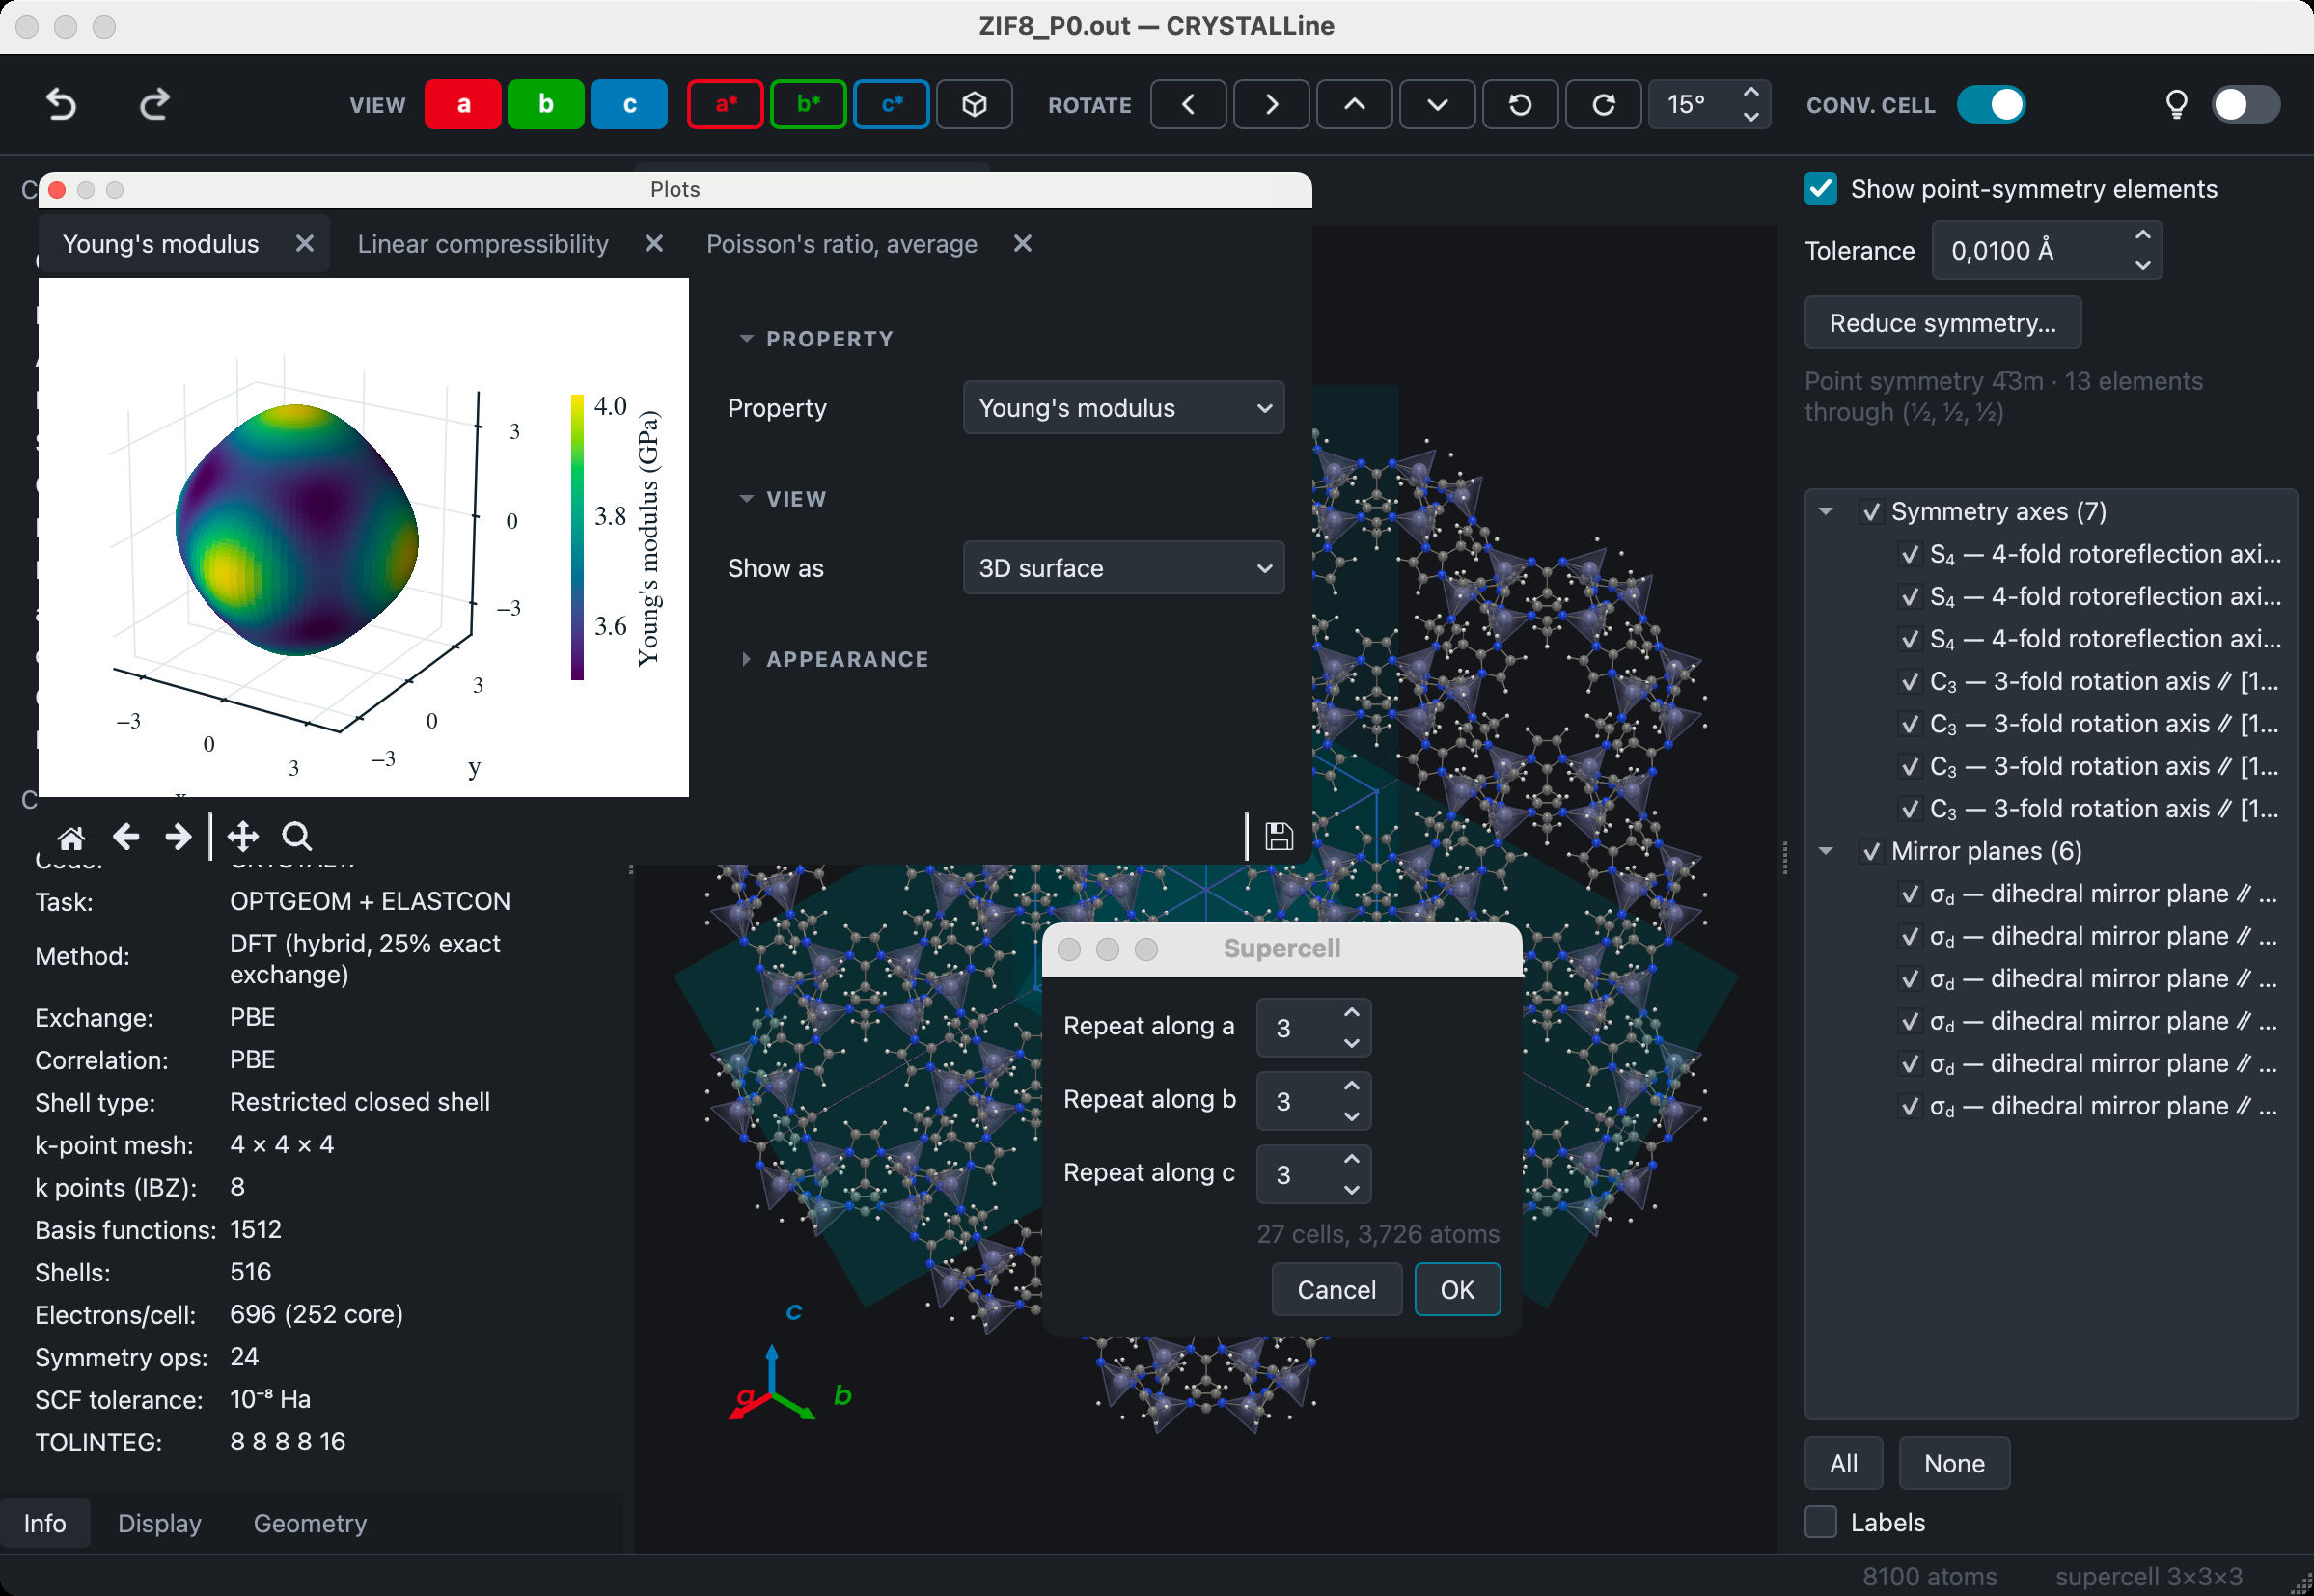
    Dear CRYSTAL community, We would like to introduce the beta release of CRYSTALLine, a new desktop app to display and edit structures for CRYSTAL, to animate phonon modes, and to plot the properties a calculation returns. It's cross-platform: it runs on Linux, macOS and Windows. It is built on CRYSTALClear, and is free software under the GNU GPL v3. CRYSTALLine follows in the tradition of MOLDRAW, written by Prof. Piero Ugliengo, which served this community for over twenty years: a program on your own machine for looking at a structure and editing it. [image: screen3.png] What it does Handling of CRYSTAL outputs, .gui/.f34 and CIF files, each in a tab of its own, with crystallographic and calculation data beside it. Automated live detection of symmetry (through spglib). Easy structure editing through drag-and-drop: add, move, delete or change atoms, build supercells, switch between primitive and crystallographic settings. Animation of vibrational modes of a frequency calculation, including out-of-Γ phonons with several exporting options. Input builder for CRYSTAL (.d12) and PROPERTIES (.d3) from the structure on screen. First Brillouin Zone visualiser and k-path selection. Advanced plotting tools (from CRYSTALClear): electronic and phonon band structure, densities of states, IR and Raman spectra (harmonic and anharmonic), elastic surfaces and sections, equations of state, and simulated XRD patterns. Most plots have their options beside them and are redrawn as these are changed. Visualisation of crystalline orbitals, charge density and electrostatic potential over the structure as isosurfaces or as slices along a lattice plane. Installing It needs Python 3.11 or newer, best in an environment of its own: conda create --name crystal python=3.12 conda activate crystal pip install CRYSTALLine crystalline The documentation has a small MgO example to try it, with a band structure and a phonon dispersion. This is a beta Accidents may take place. We would be grateful for reports of anything that fails or looks wrong, either in reply to this post or at github.com/crystaldevs/CRYSTALLine/issues. Suggestions for what to add next are more than welcome too. Links Documentation: crystaldevs.github.io/CRYSTALLine Source code: github.com/crystaldevs/CRYSTALLine PyPI: pypi.org/project/CRYSTALLine Give it a try!
  • Communications for the community and updates on upcoming events

    8 10
    8 Topics
    10 Posts
    GiacomoAmbrogio
    The Department of Chemistry and the Thomas Young Centre at Imperial College London and the Theoretical Chemistry Group of the University of Torino , in collaboration with the Computational Materials Science Group of the Science and Technology Facilities Council (STFC), are organising the [image: logo0.png] MSSC2026 Summer School on Ab initio Modelling in Solid State Chemistry The School is designed for Master and Ph.D. students, as well as for post-docs and researchers who have an interest in getting or strengthening a background in: ️ Computational Solid State Chemistry 🧠 Physics 🧱 Materials Science Surface- and Nano-Science The week-long School consists of morning lectures and afternoon hands-on tutorial sessions, where the formal framework and functionalities of the CRYSTAL electronic structure package will be explored. While we strongly encourage in-person participation, we also offer the possibility to attend remotely through streaming of morning lectures and afternoon hands-on tutorials. Participants will have the opportunity to present their research at a poster session. You can register here! Friday 8 May - Deadline for payment of early bird fees. See the School website for further details.
Suggested Topics

  • Forum Rules

    Welcome to the official forum for CRYSTAL software users! This is a space to share knowledge, find support, and connect with others interested in solid-state simulations. To maintain a productive and respectful environment, we ask all members to adhere to the following rules...

  • CRYSTALClear

    CRYSTALClear is an open source project that provides an easy Python interface with CRYSTAL. The package allows you to quickly extract information from the CRYSTAL output files and to easily generate customizable plots...


Top Users

1

Online

211

Users

159

Topics

671

Posts