fractional coordinate entry
-

Hi all, I am not getting the right geometry inputs for 1/3 and 2/3 positions. For example, for the structure below, I am trying to enter for the group 164:
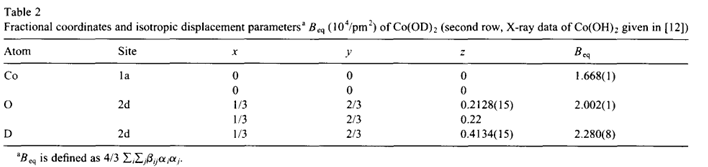
CRYSTAL 0 0 0 164 3.186 4.653 3 27 0.0 0.0 0.0 1 0.333 0.6666 0.4134 8 0.3333 0.666 0.2128 and it ends up being incorrect stoichiometry, which tells me these fractional positions were not parsed correctly. CRYSTAL CALCULATION (INPUT ACCORDING TO THE INTERNATIONAL TABLES FOR X-RAY CRYSTALLOGRAPHY) CRYSTAL FAMILY : HEXAGONAL CRYSTAL CLASS (GROTH - 1921) : DITRIGONAL SCALENOHEDRAL SPACE GROUP (CENTROSYMMETRIC) : P -3 M 1 LATTICE PARAMETERS (ANGSTROMS AND DEGREES) - CONVENTIONAL CELL A B C ALPHA BETA GAMMA 3.18600 3.18600 4.65300 90.00000 90.00000 120.00000 NUMBER OF IRREDUCIBLE ATOMS IN THE CONVENTIONAL CELL: 3 INPUT COORDINATES ATOM AT. N. COORDINATES 1 27 0.000000000000E+00 0.000000000000E+00 0.000000000000E+00 2 1 3.330000000000E-01 6.666000000000E-01 4.134000000000E-01 3 8 3.333000000000E-01 6.660000000000E-01 2.128000000000E-01 ******************************************************************************* << INFORMATION >>: FROM NOW ON, ALL COORDINATES REFER TO THE PRIMITIVE CELL ******************************************************************************* LATTICE PARAMETERS (ANGSTROMS AND DEGREES) - PRIMITIVE CELL A B C ALPHA BETA GAMMA VOLUME 3.18600 3.18600 4.65300 90.00000 90.00000 120.00000 40.903006 COORDINATES OF THE EQUIVALENT ATOMS (FRACTIONAL UNITS) N. ATOM EQUIV AT. N. X Y Z 1 1 1 27 CO 0.00000000000E+00 0.00000000000E+00 0.00000000000E+00 2 2 1 1 H 3.33000000000E-01 -3.33400000000E-01 4.13400000000E-01 3 2 2 1 H 3.33400000000E-01 -3.33600000000E-01 4.13400000000E-01 4 2 3 1 H 3.33600000000E-01 -3.33000000000E-01 4.13400000000E-01 5 2 4 1 H -3.33600000000E-01 3.33400000000E-01 -4.13400000000E-01 6 2 5 1 H -3.33000000000E-01 3.33600000000E-01 -4.13400000000E-01 7 2 6 1 H -3.33400000000E-01 3.33000000000E-01 -4.13400000000E-01 8 2 7 1 H -3.33000000000E-01 3.33400000000E-01 -4.13400000000E-01 9 2 8 1 H -3.33400000000E-01 3.33600000000E-01 -4.13400000000E-01 10 2 9 1 H -3.33600000000E-01 3.33000000000E-01 -4.13400000000E-01 11 2 10 1 H 3.33600000000E-01 -3.33400000000E-01 4.13400000000E-01 12 2 11 1 H 3.33000000000E-01 -3.33600000000E-01 4.13400000000E-01 13 2 12 1 H 3.33400000000E-01 -3.33000000000E-01 4.13400000000E-01 14 3 1 8 O 3.33300000000E-01 -3.34000000000E-01 2.12800000000E-01 15 3 2 8 O 3.34000000000E-01 -3.32700000000E-01 2.12800000000E-01 16 3 3 8 O 3.32700000000E-01 -3.33300000000E-01 2.12800000000E-01 17 3 4 8 O -3.32700000000E-01 3.34000000000E-01 -2.12800000000E-01 18 3 5 8 O -3.33300000000E-01 3.32700000000E-01 -2.12800000000E-01 19 3 6 8 O -3.34000000000E-01 3.33300000000E-01 -2.12800000000E-01 20 3 7 8 O -3.33300000000E-01 3.34000000000E-01 -2.12800000000E-01 21 3 8 8 O -3.34000000000E-01 3.32700000000E-01 -2.12800000000E-01 22 3 9 8 O -3.32700000000E-01 3.33300000000E-01 -2.12800000000E-01 23 3 10 8 O 3.32700000000E-01 -3.34000000000E-01 2.12800000000E-01 24 3 11 8 O 3.33300000000E-01 -3.32700000000E-01 2.12800000000E-01 25 3 12 8 O 3.34000000000E-01 -3.33300000000E-01 2.12800000000E-01 -
Hi,
please try to define the atomic positions with more figures.
This is important for 1/3 and 2/3 fractional coordinates.CRYSTAL 0 0 0 164 3.186 4.653 3 27 0.0 0.0 0.0 1 0.33333333 0.66666666 0.4134 8 0.33333333 0.66666666 0.2128I hope it helps.
Giuseppe

Hello! It looks like you're interested in this conversation, but you don't have an account yet.
Getting fed up of having to scroll through the same posts each visit? When you register for an account, you'll always come back to exactly where you were before, and choose to be notified of new replies (either via email, or push notification). You'll also be able to save bookmarks and upvote posts to show your appreciation to other community members.
With your input, this post could be even better 💗
Register Login