Skip to content

Other Questions

8 Topics 27 Posts

Questions that do not fit in other categories

  • Creating Wannierization Input Files

    3
    0 Votes
    3 Posts
    537 Views
    CaesarSalad
    Hi Giacomo Now that you pointed that out... I can see that my launch script was at fault indeed. I wasn't actually passing the correct file name to the launch script. It is running well now, thanks a lot! C.
  • Localized Crystalline Orbitals

    5
    1
    0 Votes
    5 Posts
    697 Views
    Chhatra
    Thankyou so much for your guidance. I will try!
  • Troubles with coordinate system in output using POTC

    3
    0 Votes
    3 Posts
    372 Views
    denkos
    Okay, so then I need to sort the values manually according to convention. Thank you!
  • Problem with COs: Crystalline Orbitals

    3
    0 Votes
    3 Posts
    372 Views
    yuliyaglushkova
    Thank you!
  • Query regarding COHP Calculation

    3
    2
    0 Votes
    3 Posts
    787 Views
    Rams
    Hi Prof. Erba, Thanks for the reply. From the file, I could see there are value from energy " -7.9324E-01" (Line No: 81826). I could plot the data using Python, Xmgrace. With CRYSPLOT somehow the data loading is slow and my browser hangs so I am not able to check it. But definitely there is data. I requested for COHP between Ni and three nearby oxygen. In the file there are 5 columns, so if I am not mistaken it corresponds to "Energy - Next three columns in the order I request - Final Column of DOS ?" (Because the last column data when plotted matches the DOS) Thanks & Regards, Rams
  • problem with COOP calculation

    3
    0 Votes
    3 Posts
    625 Views
    mriveraa
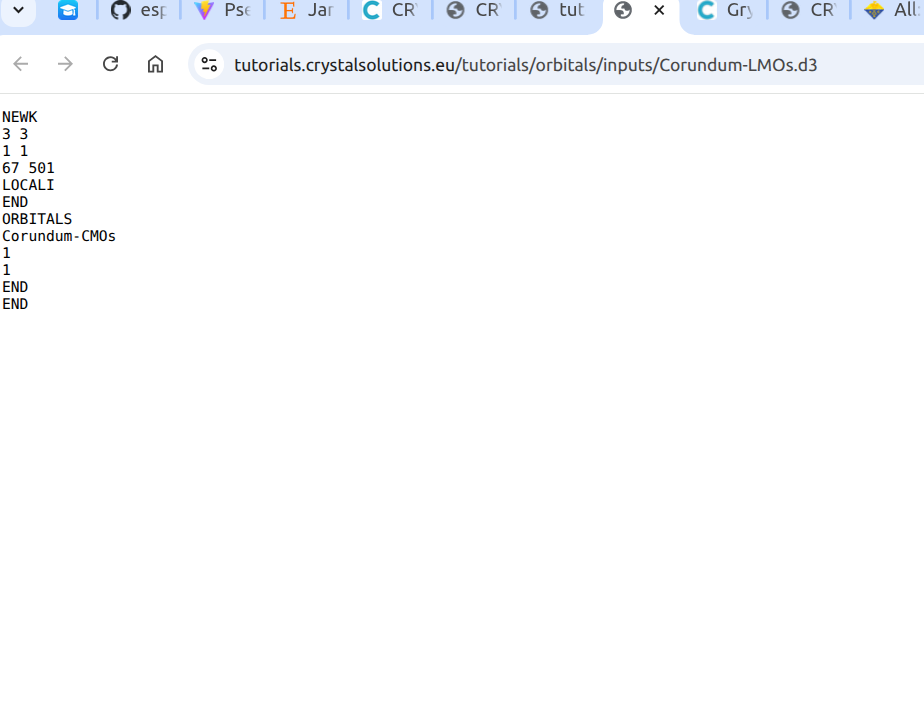
    Hi! In a COOP calculation you aim to have a description on the interaction between pairs of orbitals or atoms. In order to do that, you need to indicate each pair you are interested to analyze. In your input, in the first line after the COOP keyword, the initial number 1 indicates that you are interested in one pair of orbitals/atoms. You still need to indicate a pair of orbitals or atoms to be considered, writing them in separated lines. Consider this example, taken from the Tutorials webpage: NEWK 6 6 1 0 COOP 1 200 7 14 1 12 0 -1 1 -1 2 END Here, the two lines before the final END keyword indicate which atoms will be considered (atoms, given that the lines start with a negative value, as stated in the manual page 322). COOP will be evaluated considering the first and second atoms of the systems (with indices 1 and 2). From your previous calculations you can recover the indices of the atoms/orbitals you are interested. Let me know if this information has been useful
  • Anisotropic net in properties

    2
    0 Votes
    2 Posts
    507 Views
    eascrizzi
    Hi aimipa! Yes there is an option to set an anisotropic grid from NEWK, you can find it at page 346 of the CRYSTAL User's Manual, see also the screenshot I attach below. You have to set IS=0 and then you can choose three different shrinking factors along B1, B2 and B3. For example: NEWK 0 12 12 8 4 sets a shrinking factor of 12, 8 and 4 along the three reciprocal lattice vectors. I have never tested this anisotropic option in a BOLTZTRA calculation, would you mind let me know if it works? [image: newk.png]
  • Optical Properties calculation using CRYSTAL code

    5
    0 Votes
    5 Posts
    1k Views
    R.Zosiamliana
    Thank you Jacques for your kind help. This will solve multiple problems of my current work Regards R.Zosiamliana