P 21/a symmetry not found
-

I am trying to input the following cif file Mg, Acta Crystallographica 17 (1964) 1478-1479.cif which is P 21/a space group 14. However, in the manual space group 14 is P 21/c and my geometry ends up looking funky. I also tried entering manually, e.g. P 2 1a but that simply does not work. Can you help me enter the correct space group here?
CRYSTAL
1 0 0
P 2 1a
9.383 12.669 6.220 107.05
20
7 0.1321 0.3509 0.3611
1 0.058 0.337 0.225
1 0.208 0.305 0.394
1 0.095 0.344 0.487
1 0.174 0.421 0.346
12 0 0 0
8 0.1603 -0.1094 -0.0307
8 0.1685 0.1042 0.1656
8 -0.0017 -0.0687 0.2986
1 0.2 0.091 0.317
1 0.227 0.134 0.116
1 0.252 -0.096 0.059
1 0.143 -0.176 -0.008
1 -0.097 -0.066 0.341
1 0.027 -0.135 0.325
16 0.0953 -0.3605 0.2575
8 -0.0469 -0.4174 0.2116
8 0.2185 -0.4328 0.3718
8 0.1185 -0.3211 0.0456
8 0.0951 -0.2702 0.4089 -
I think I got it but I do not know how to explain it. I set it up as below after lots of trial and error
MgNH4SO4*6H2O from Mg, Acta Crystallographica 17 (1964) 1478-1479
CRYSTAL
1 0 0
P 1 21/a 1
9.383 12.669 6.220 107.05
20
7 0.1321 0.3509 0.3611
1 0.058 0.337 0.225
1 0.208 0.305 0.394
1 0.095 0.344 0.487
1 0.174 0.421 0.346
12 0 0 0
8 0.1603 -0.1094 -0.0307
8 0.1685 0.1042 0.1656
8 -0.0017 -0.0687 0.2986
1 0.2 0.091 0.317
1 0.227 0.134 0.116
1 0.252 -0.096 0.059
1 0.143 -0.176 -0.008
1 -0.097 -0.066 0.341
1 0.027 -0.135 0.325
16 0.0953 -0.3605 0.2575
8 -0.0469 -0.4174 0.2116
8 0.2185 -0.4328 0.3718
8 0.1185 -0.3211 0.0456
8 0.0951 -0.2702 0.4089
EOS
RANGE
0.94 1.06 8
PREOPTGEOM
MAXCYCLE
500
END
BASISSET
pob-TZVP-rev2
DFT
B3LYP-D3
XLGRID
END
TOLINTEG
8 8 8 8 16
SHRINK
4 4
BIPOSIZE
41202400
EXCHSIZE
41202400
END -

Hi,
I think it was just a syntax problem on the way the symbol of the space group was inputted. This is how it should look like in this case:
TEST
CRYSTAL
1 0 0
P 21/A
9.383 12.669 6.220 107.05
20
7 0.1321 0.3509 0.3611
1 0.058 0.337 0.225
1 0.208 0.305 0.394
1 0.095 0.344 0.487
1 0.174 0.421 0.346
12 0 0 0
8 0.1603 -0.1094 -0.0307
8 0.1685 0.1042 0.1656
8 -0.0017 -0.0687 0.2986
1 0.2 0.091 0.317
1 0.227 0.134 0.116
1 0.252 -0.096 0.059
1 0.143 -0.176 -0.008
1 -0.097 -0.066 0.341
1 0.027 -0.135 0.325
16 0.0953 -0.3605 0.2575
8 -0.0469 -0.4174 0.2116
8 0.2185 -0.4328 0.3718
8 0.1185 -0.3211 0.0456
8 0.0951 -0.2702 0.4089
CIFPRT
TESTGEOM
END
END
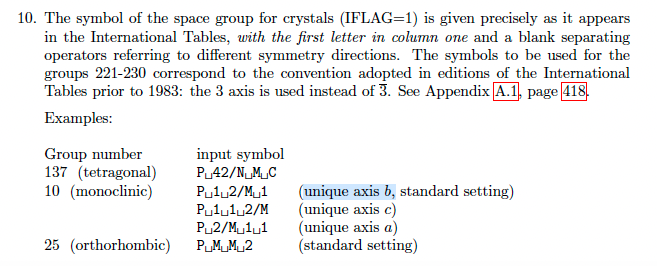
For future reference, the syntax for space group symbols in CRYSTAL is discussed at pages 22-23 of the CRYSTAL23 User's Manual (see screenshot below):
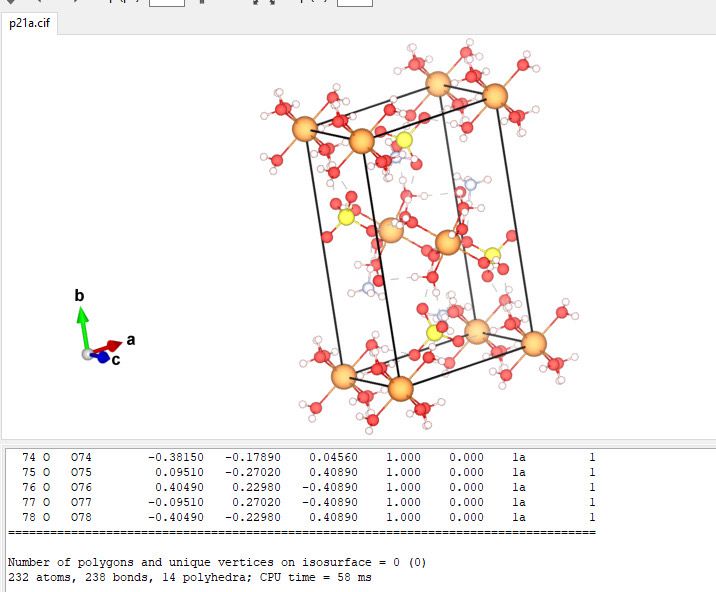
We ran a test and the structure built from CRYSTAL looks reasonable:
Hope this helps,
Alessandro Erba
Professor of Physical Chemistry
Department of Chemistry, University of Torino
[email protected]

Hello! It looks like you're interested in this conversation, but you don't have an account yet.
Getting fed up of having to scroll through the same posts each visit? When you register for an account, you'll always come back to exactly where you were before, and choose to be notified of new replies (either via email, or push notification). You'll also be able to save bookmarks and upvote posts to show your appreciation to other community members.
With your input, this post could be even better 💗
Register Login