Skip to content
  • Home
  • Recent
Collapse
Brand Logo
CRYSTAL23
Latest v1.0.1
Tutorials Try the Demo Get a License
Tutorials Try the Demo Get a License Instagram
aerba

Alessandro Erba

@aerba
Developer
Unfollow Follow
About
Posts
115
Topics
1
Shares
0
Groups
1
Followers
9
Following
8

Posts

Recent Best Controversial

  • corrupted size vs. prev_size while consolidating
    aerba aerba
    Bug Reports

    Hi,

    After running a few tests, we found a bug towards the end of the EOS module due to the size mismatch of different arrays (specifically, in the full list of direct lattice vectors of the equilibrium configuration and maximally compressed configuration). While this requires an actual fix to the code, we also found a workaround solution for you. Indeed, if the maximally compressed configuration has the same list as for the equilibrium configuration, everything is fine. Thus, in your case, it was enough to explore a maximum compression of 5% (rather than 8%) to be able to run smoothly the job.

    Please, find here a working input file (where we have also reduced the number of explored volumes to 4+1 to speed up the calculation without resulting in any significant loss of accuracy).


  • corrupted size vs. prev_size while consolidating
    aerba aerba
    Bug Reports

    job314 I wouldn't be THAT optimistic 😉


  • MSSC2025 Summer School, 8-12 September 2025, Torino (Italy) "Ab initio Modelling in Solid State Chemistry"
    aerba aerba
    Events

    Dear CRYSTALlers,

    The Theoretical Chemistry Group of the University of Torino and the Department of Chemistry and the Thomas Young Centre at Imperial College London are organising the

    MSSC2025 Summer School on Ab initio Modelling in Solid State Chemistry

    logo_Full.png

    to be held in Torino (Italy), 8-12 September 2025. The School is designed for Master and Ph.D. students, as well as for post-docs and researchers who have an interest in getting or strengthening a background in Computational Solid State Chemistry, Physics, Materials Science, Surface- and Nano-Science.

    The week-long School consists of morning lectures and afternoon hands-on tutorial sessions, where the formal framework and functionalities of the CRYSTAL electronic structure package will be explored.

    Registration opens on March 15 2025. See the School website for further details.

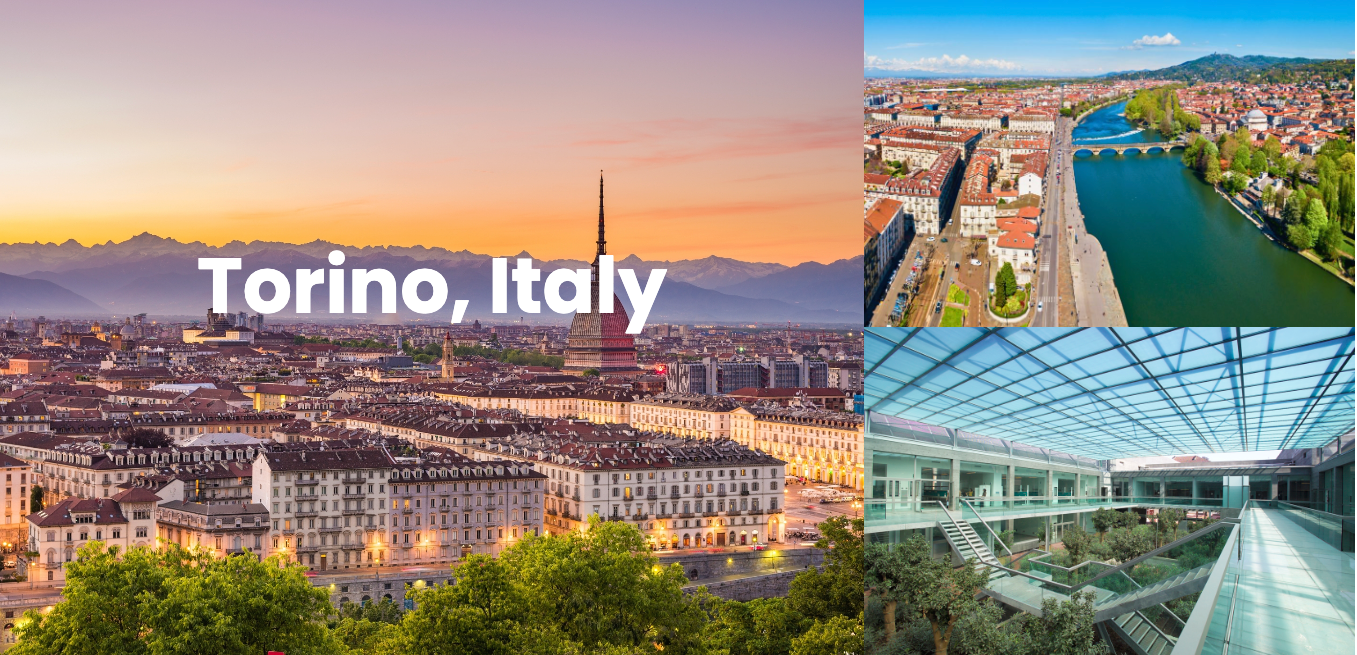
    The School takes place in the elegant city of Torino (Italy):

    Torino.png

    ­
    While we strongly encourage in-person participation, we also offer the possibility to attend remotely through streaming of morning lectures and afternoon hands-on tutorials.

    Participants will have the opportunity to present their research at a poster session.

    See you in Torino!


  • exporting optaxxx to cif
    aerba aerba
    Others

    job314 Yes, it should print the geometry of the last optimization step. Please, be aware that the CIFPRT and CIFPRTSYM options are not as general as one would like them to be. For systems where the primitive and crystallographic cells differ, they may result in an incomplete list of atoms. In those cases, I recommend switching symmetry off with the SYMMREMO option in combination with TESTGEOM, just to generate the .cif files correctly.


  • Confusion in ELASTCON output:
    aerba aerba
    Elasticity, Piezoelectricity, Photoelasticity

    Hi,

    Elastic constants of a 3D lattice in CRYSTAL are defined and computed as:

    $$
    C_{ij}^\textup{3D} = \frac{1}{V} \left( \frac{\partial^2 E}{\partial \eta_i \partial \eta_j}\right)
    $$

    where the strain \( \eta \) is dimensionless and thus the elastic constants have units of \( \textup{energy/length}^3 \), which corresponds to force/surface (i.e. a pressure).

    For 1D and 2D periodic lattices, as the volume \(V\) is not defined, one may divide by the length \(l \) and area \( A\) of the lattice cell instead:

    $$
    C_{ij}^\textup{1D} = \frac{1}{l} \left( \frac{\partial^2 E}{\partial \eta_i \partial \eta_j}\right) \qquad \textup{and} \qquad C_{ij}^\textup{2D} = \frac{1}{A} \left( \frac{\partial^2 E}{\partial \eta_i \partial \eta_j}\right)
    $$

    However, in CRYSTAL for 1D and 2D lattices we do not divide by \(l \) or \( A\) , and just define and compute the elastic constants as:

    $$
    C_{ij}^\textup{1D and 2D} = \left( \frac{\partial^2 E}{\partial \eta_i \partial \eta_j}\right)
    $$

    with units of energy. So, starting from the constants printed in the CRYSTAL output all you have to do is divide by the area of the 2D cell if you need them expressed in \( \textup{energy/length}^2 \).


  • corrupted size vs. prev_size while consolidating
    aerba aerba
    Bug Reports

    We are running some tests and I think we are close to identifying the problem. We’ll probably have a solution by the beginning of next week.


  • cam-B3LYP with pobTZVP SCF convergence
    aerba aerba
    Single-Point Calculations

    job314 Good point! About the shrinking factor: the anisotropic shrinking factor in CRYSTAL does not work properly for those calculations where the symmetry of the system may change (for instance in frequency calculations, FREQCALC, where displaced nuclear configurations are explored, or elastic calculations, ELASTCON, where the lattice is strained, etc.). So in general, I personally tend to avoid using an anisotropic shrinking factor.

    However, for symmetry-preserving calculations (such as SCF, OPTGEOM, EOS) the use of an anisotropic shrinking factor should be fine.


  • MAXCYCLE is not functional within EOS and PREOPTGEOM
    aerba aerba
    Equation-of-State and Pressure

    job314 I have checked your attached output files. The OPTGEOM calculation converges to the optimized structure in 29 steps. The PREOPTGEOM calculation within the EOS option converges to the same optimized structure in 25 steps. A lower number of steps in EOS is likely due to the higher accuracy of the computed forces because of tighter default settings in EOS than in OPTGEOM (for instance, TOLDEE is set to 8 in EOS and to 7 in OPTGEOM).

    The EOS.out file you shared looks perfectly fine. I can not find any failed geometry optimizations there. After the pre-optimization, the actual constant-volume optimizations are performed (converged in 40, 19, 16, 16, 8 steps, respectively). The output is not complete because I guess the calculation was still running.


  • cam-B3LYP with pobTZVP SCF convergence
    aerba aerba
    Single-Point Calculations

    Hi,

    Find below a modified input file for which the SCF converges in 25 iterations:

    Na2Si2O5, Rakic Physics and Chemistry of Minerals 29 (2002) 477-484 - Ale mod
    CRYSTAL
    0 0 0
    14
    4.725582 23.296569 7.936255 90.15
    18
    11 .2299 .26138 .5247
    11 .2743 .51408 .3548
    11 .2534 .47281 -.1019
    11 .7548 .28108 .2783
    14 .2961 .36325 .1966
    14 .6843 .34077 .6429
    14 .1831 .40821 .5373
    14 .7948 .38941 -.0185
    8 .1210 .3844 .0352
    8 .2339 .3015 .2523
    8 .6209 .3690 .1429
    8 .2440 .4105 .3392
    8 .7431 .3414 -.1590
    8 .7456 .2819 .5655
    8 .3573 .3561 .6162
    8 -.1423 .3920 .5597
    8 .2452 .4667 .6166
    8 .7336 .4511 -.0767
    EOS
    RANGE
    0.95 1.05 8
    PREOPTGEOM
    MAXCYCLE
    500
    END
    BASISSET
    POB-TZVP-REV2
    DFT
    cam-B3LYP
    XLGRID
    END
    TOLINTEG
    10 10 10 10 20
    SHRINK
    6 6
    BIPOSIZE
    11202400
    EXCHSIZE
    11202400
    MAXCYCLE
    200
    END
    

    We have increased the values of TOLINTEG, used an isotropic shrinking factor, and re-activated the DIIS accelerator.


  • MAXCYCLE is not functional within EOS and PREOPTGEOM
    aerba aerba
    Equation-of-State and Pressure

    job314 I see, interesting! I don't seem to be able to find the input/output files


  • MAXCYCLE is not functional within EOS and PREOPTGEOM
    aerba aerba
    Equation-of-State and Pressure

    Yes, the pre-optimization of the EOS is indeed a full structural relaxation where cell/volume/atomic positions are all relaxed at once. If you want me to have a closer look at this case, please send me the input/output files.


  • MAXCYCLE is not functional within EOS and PREOPTGEOM
    aerba aerba
    Equation-of-State and Pressure

    Hi,

    Yes, it looks strange, and 102 is a funny number.

    Could you try to run a regular optimization (i.e. with OPTGEOM and not within the EOS) and let me know if the same happens?


  • computing anharmonic shifts for C-H vibrations
    aerba aerba
    Harmonic and Anharmonic Lattice Dynamics and Thermodynamics

    Hi,

    In CRYSTAL23, the anharmonicity of a given vibration mode - or of a set thereof - can be computed via: I) the evaluation of cubic and quartic interatomic force constants (in the basis of the normal modes) followed by II) the solution of the nuclear Schroedinger equation with either the vibrational self-consistent field (VSCF) or vibrational configuration interaction (VCI) method.

    Details on the actual implementation of steps I) and II) can be found here:

    I) Anharmonic force constants

    II) VSCF and VCI for solids

    Step-by-step, the procedure is as follows:

    1. Geometry optimization to fully relax atomic positions within the cell (OPTGEOM keyword);
    2. Harmonic frequency calculation (FREQCALC keyword);
    3. Selection of the normal modes for which the anharmonic correction is to be computed (for instance, in your case, those corresponding to C-H stretching vibrations);
    4. Calculation of cubic and quartic interatomic force constants for the selected modes + VSCF (or VCI) calculation of anharmonic states.

    As an example, let' s assume that C-H stretching vibrations correspond to modes 15-20 in the list generated from the harmonic calculation (that is, there are 6 different C-H stretching modes). The input for steps 3. and 4. above would read:

    FREQCALC
    RESTART
    ANHAPES
    6
    15 16 17 18 19 20
    3 0.9
    VSCF
    END
    

    that is, we restart the harmonic frequency calculation, and where 6 is the number of modes, 15 16 17 18 19 20 are the selected modes, and 3 0.9 are two parameters specifying the numerical approach used for the evaluation of cubic and quartic force constants.

    Please, note that with such a calculation not only the "intrinsic" anharmonicity of each selected mode is evaluated but also the couplings among all selected modes. If, instead, one just wants to compute the "intrinsic" anharmonicity with no couplings, independent calculations can be run, one per each selected mode.

    Visit this page for a tutorial on anharmonic calculations in CRYSTAL

  • Imaginary frequencies
    aerba aerba
    Vibrational Spectroscopies: IR, Raman, INS

    Hi,

    The presence of imaginary frequencies is a sign that the geometry is not a minimum of the potential energy surface (PES). In general, this may due to two main factors:

    1) A somewhat loose overall numerical precision in the geometry optimization + harmonic frequencies calculation. Here, it seems that you have already explored a few parameters. We can distinguish between parameters governing the overall numerical precision of the SCF + forces calculations, and those that are specific to the evaluation of the Hessian:

    1.1) Precision of SCF+forces

    One may increase a bit the thresholds for the screening of two-electron integrals (TOLINTEG keyword), switch-off the bipolar approximation (NOBIPOLA keyword), increase the shrinking factor (SHRINK keyword), use a denser grid for numerical integration of the exchange-correlation term (see XLGRID, XXLGRID keywords), tighten the convergence criteria for the SCF (setting TOLDEE to 10 or 11 for instance), tighten the convergence criteria for the geometry optimization step (see TOLDEG and TOLDEX keywords).

    1.2) Numerical evaluation of the Hessian

    In many cases, a more numerically stable evaluation of the Hessian is achieved by use of a two-sided finite difference approach (rather than the default one-sided approach). This can be activated with the NUMDERIV keyword within the FREQCALC input block as follows:

    FREQCALC
    NUMDERIV
    2
    ENDFREQ
    

    2) The presence of symmetry-constraints that prevent the optimizer to get to the minimum of the PES. If this is the case, removing symmetry constraints may be key to reach the minimum. This can be done by use of the SYMMREMO keyword (to be inserted in the geometry input block).


  • IR and Raman frequency calculations at different temperature
    aerba aerba
    Vibrational Spectroscopies: IR, Raman, INS

    Hi,

    By default, thermodynamic properties on top of harmonic frequencies are computed at room temperature. Different values of temperature can be explored by use of the TEMPERAT keyword. To restart the harmonic frequency calculation, use the RESTART keyword. An example is given below:

    FREQCALC
    RESTART
    TEMPERAT
    5 200 600
    END
    

    With the input above, thermodynamic properties will be computed and printed at 5 temperatures, equally spaced in the range 200-600 K.

  • 1 / 1
  • Login

  • Don't have an account? Register

  • Login or register to search.
  • First post
    Last post
0
  • Home
  • Recent