Skip to content
  • How create band structure in P1 symmetry

    Band Structure
    3
    0 Votes
    3 Posts
    667 Views
    aerba
    Hi, Let me just add that of course in P1 there are no special high-symmetry points to guide you in the definition of the path, so you need to be a little creative. For instance, you can start from Gamma (0 0 0) and go to the edge of the FBZ along the b1 reciprocal lattice (1/2 0 0), to then go to (1/2 1/2 0), then to (0 1/2 0) then back to Gamma (0 0 0) and then to the edge along the b3 reciprocal lattice (0 0 1/2). Or something else!
  • segmentation fault with double free or corruption during SCF

    Bug Reports
    6
    0 Votes
    6 Posts
    1k Views
    GiacomoAmbrogio
    Hi andrejsc, There is no "hardcoded" switch in the TOLINTEG keyword, I suspect that the quote from the manual comes from the "old days", when a threshold of \( 10^{-20} \) was considered absurdly small. However, with advances in hardware and the increasingly complex structures we want to compute, such thresholds are sometimes necessary. A small note: don't warry about going past machine precision with these small numbers. All evaluations of integral thresholds in the code are performed at the logaritmic level (ie only on the exponent), so you should be fine even with a value of 1 million in TOLINTEG (though maybe not fine in terms of computation time)
  • D infinity h symmetry

    Geometry Editing
    4
    0 Votes
    4 Posts
    793 Views
    dmitoli
    If you fear that a reduced symmetry might influence the thermodynamic results, you can simply run the calculation without imposing any symmetry at all. For a 3-atom system like CO₂ the computational cost is so minimal that you can run both symmetry-free and \(D_{4h}\) calculations. Comparing the two results should give you an indication of whether the imposed symmetry has any impact on the quantities you are interested in.
  • PDOS Projection for Individual Orbital Components

    Density-of-States
    2
    0 Votes
    2 Posts
    599 Views
    GiacomoAmbrogio
    Hi Orion, Looking at your input, I think you want to plot the orbital projected electronic DOS (in CRYSTAL this is done with the DOSS keyword, PDOS in CRYSTAL normally refers to phonon DOS). To obtain separate px py pz projections, the correct approach is: set the first parameter of DOSS to 2 (because you want two separate projections). After the main DOSS line, you must then add one line per projection, specifying which atomic orbitals (AOs) belong to each projection. Each line has the form: n AOs index_1 index_2 ... index_n The AO indices correspond to the ones printed in the CRYSTAL output. To find them, search in the SCF output for "LOCAL ATOMIC FUNCTIONS BASIS SET", you will see a table with all the basis set, that will look like this: ******************************************************************************* ATOM X(AU) Y(AU) Z(AU) N. TYPE EXPONENT S COEF P COEF D/F/G COEF ******************************************************************************* 1 O -2.793 -4.838 -4.110 1 S 8.589E+03 1.895E-03 0.000E+00 0.000E+00 1.297E+03 1.439E-02 0.000E+00 0.000E+00 2.993E+02 7.073E-02 0.000E+00 0.000E+00 8.738E+01 2.400E-01 0.000E+00 0.000E+00 2.568E+01 5.948E-01 0.000E+00 0.000E+00 3.740E+00 2.808E-01 0.000E+00 0.000E+00 2- 5 SP 4.212E+01 1.139E-01 3.651E-02 0.000E+00 9.628E+00 9.208E-01 2.372E-01 0.000E+00 2.853E+00-3.274E-03 8.197E-01 0.000E+00 6- 9 SP 9.057E-01 1.000E+00 1.000E+00 0.000E+00 10- 13 SP 2.556E-01 1.000E+00 1.000E+00 0.000E+00 14- 18 D 1.292E+00 0.000E+00 0.000E+00 1.000E+00 The indeces are the numbers befor each orbital shell (1 2-5 6-9 10-13 14-18). The notation 2-5 means indices 2 through 5, because that shell contains 4 AOs. From these blocks you must identify the p-type orbitals for each atom (Zn and Cl). The p shells will appear in groups of three basis functions (px, py, pz). So, in your system: Locate the Zn atom in this printed basis list Identify the block corresponding to its p orbitals Extract the AO indices for px, py, pz Do the same for Cl Then you define one projection per line. For example, structurally something like: NEWK 12 12 1 0 DOSS 2 100 3 6 1 12 0 3 <list of Zn p AOs> 3 <list of Cl p AOs> END Keep in mind that, if your basis is double- or triple-Z, each p shell appears as a separate triplet of AOs (one triplet per Z). To correctly project all p you must include the corresponding AOs from every p-shell triplet for that atom. Let me know if you manage to do this, or if you need further help.
  • extract asymmetric fragment

    Geometry Editing
    4
    0 Votes
    4 Posts
    795 Views
    dmitoli
    Dear Jonas, I tried using pymatgen to extract the point-symmetry information from your .xyz file (see the Python script below): from pymatgen.core import Molecule from pymatgen.symmetry import analyzer bigstructure = Molecule.from_file("yourfile.xyz") PGstructure = analyzer.PointGroupAnalyzer(bigstructure) sym_mol = PGstructure.get_equivalent_atoms() print(sym_mol["eq_sets"]) This returns a Python data structure containing the symmetry-irreducible sets of atoms (only 6 for this system, which is indeed of Ih point group!). When preparing the CRYSTAL input, be careful with the orientation of your asymmetric unit. In my case, for example, I had to change the sign of the x and y coordinates to make the symmetry consistent with CRYSTAL’s conventions. Icosahedral point groups are available in CRYSTAL (Ih is point group number 47 in CRYSTAL), so the input fort this molecular cage reads: Symm. structure MOLECULE 47 6 8 2.605032231 -11.914271806 11.762689798 6 4.344538598 15.664236912 4.366798884 6 3.427479683 14.428996321 8.326818862 6 -8.906580884 1.370529673 14.411150640 1 2.632960331 14.962480562 7.832453804 5 -8.776255231 -2.916256114 14.200279302 COORPRT TESTGEOM END @aerba Christmas is already in the air indeed!
  • SCF convergence ferromagnetic MIL-127

    Single-Point Calculations
    5
    0 Votes
    5 Posts
    1k Views
    wgtemmer
    Alright, it sounds like the difficulties originate from the system itself, rather than the particular CRYSTAL input. Thanks again for all the help! Best regards, Wim
  • OPTGEOM for RUNCONFS

    Geometry Optimisations
    3
    0 Votes
    3 Posts
    756 Views
    Igor12
    @aerba, Thank you very much for your reply! Unfortunately, I don't speak English very well, which is why I may have phrased my question incorrectly. My question was that the function OPTGEOM does not work correctly with RUNCONFS. I had already tried the steps you suggested. However, I was able to figure out the problem. For geometric optimization, I just needed to specify the keyword more correctly: CRYSTAL 0 0 0 194 3.065 17.656 4 22 0 0 0.5 14 0 0 0.75 22 0.666666 0.333333 0.364919 6 0.333333 0.666666 0.427507 SCELCONF 1 0 0 0 1 0 0 0 1 RUNCONFS ATOMSUBS 22 273 OPTGEOM MAXCYCLE 500 ENDOPT END END I'll leave it here, maybe it will be useful to someone in the future. Nevertheless, thank you very much for your responsiveness
  • SCANMODE io error Read_int_1d

    Vibrational Spectroscopies: IR, Raman, INS
    8
    0 Votes
    8 Posts
    1k Views
    aerba
    Hi, @job314 said in SCANMODE io error Read_int_1d: So it turns out having fort.13 and fort.20 is not optional for restart. Yes, the lack of these two files was the origin of the I/O error. @job314 said in SCANMODE io error Read_int_1d: PS. I would also like to ask developers to allow uploading compressed files to this forum Now also .zip, .tar, .tgz, .tar.gz files can be uploaded. Cheers,
  • Double Walled Naontubes X(n1,0)@Y(n2>>n1,0)

    Geometry Optimisations
    9
    0 Votes
    9 Posts
    1k Views
    Drmajouri2025
    Good morning, OH that’s cool ️, here we go and that is exactly what I want… Thank you again Dr. Casassa
  • 0 Votes
    4 Posts
    1k Views
    similt
    Thanks. SMEAR helped me to converge scf.
  • HELP WITH COMPOSITE METHODS

    Basis Sets
    2
    0 Votes
    2 Posts
    405 Views
    bcivalleri
    Dear George, According to your output, you are using CRYSTAL17 to perform the HFsol-3c calculation. Please note that HFsol-3c and the other "sol-3c" composite methods are only available in CRYSTAL23. Best regards, Bartolomeo
  • Query regarding COHP Calculation

    Other Questions
    3
    2
    0 Votes
    3 Posts
    792 Views
    Rams
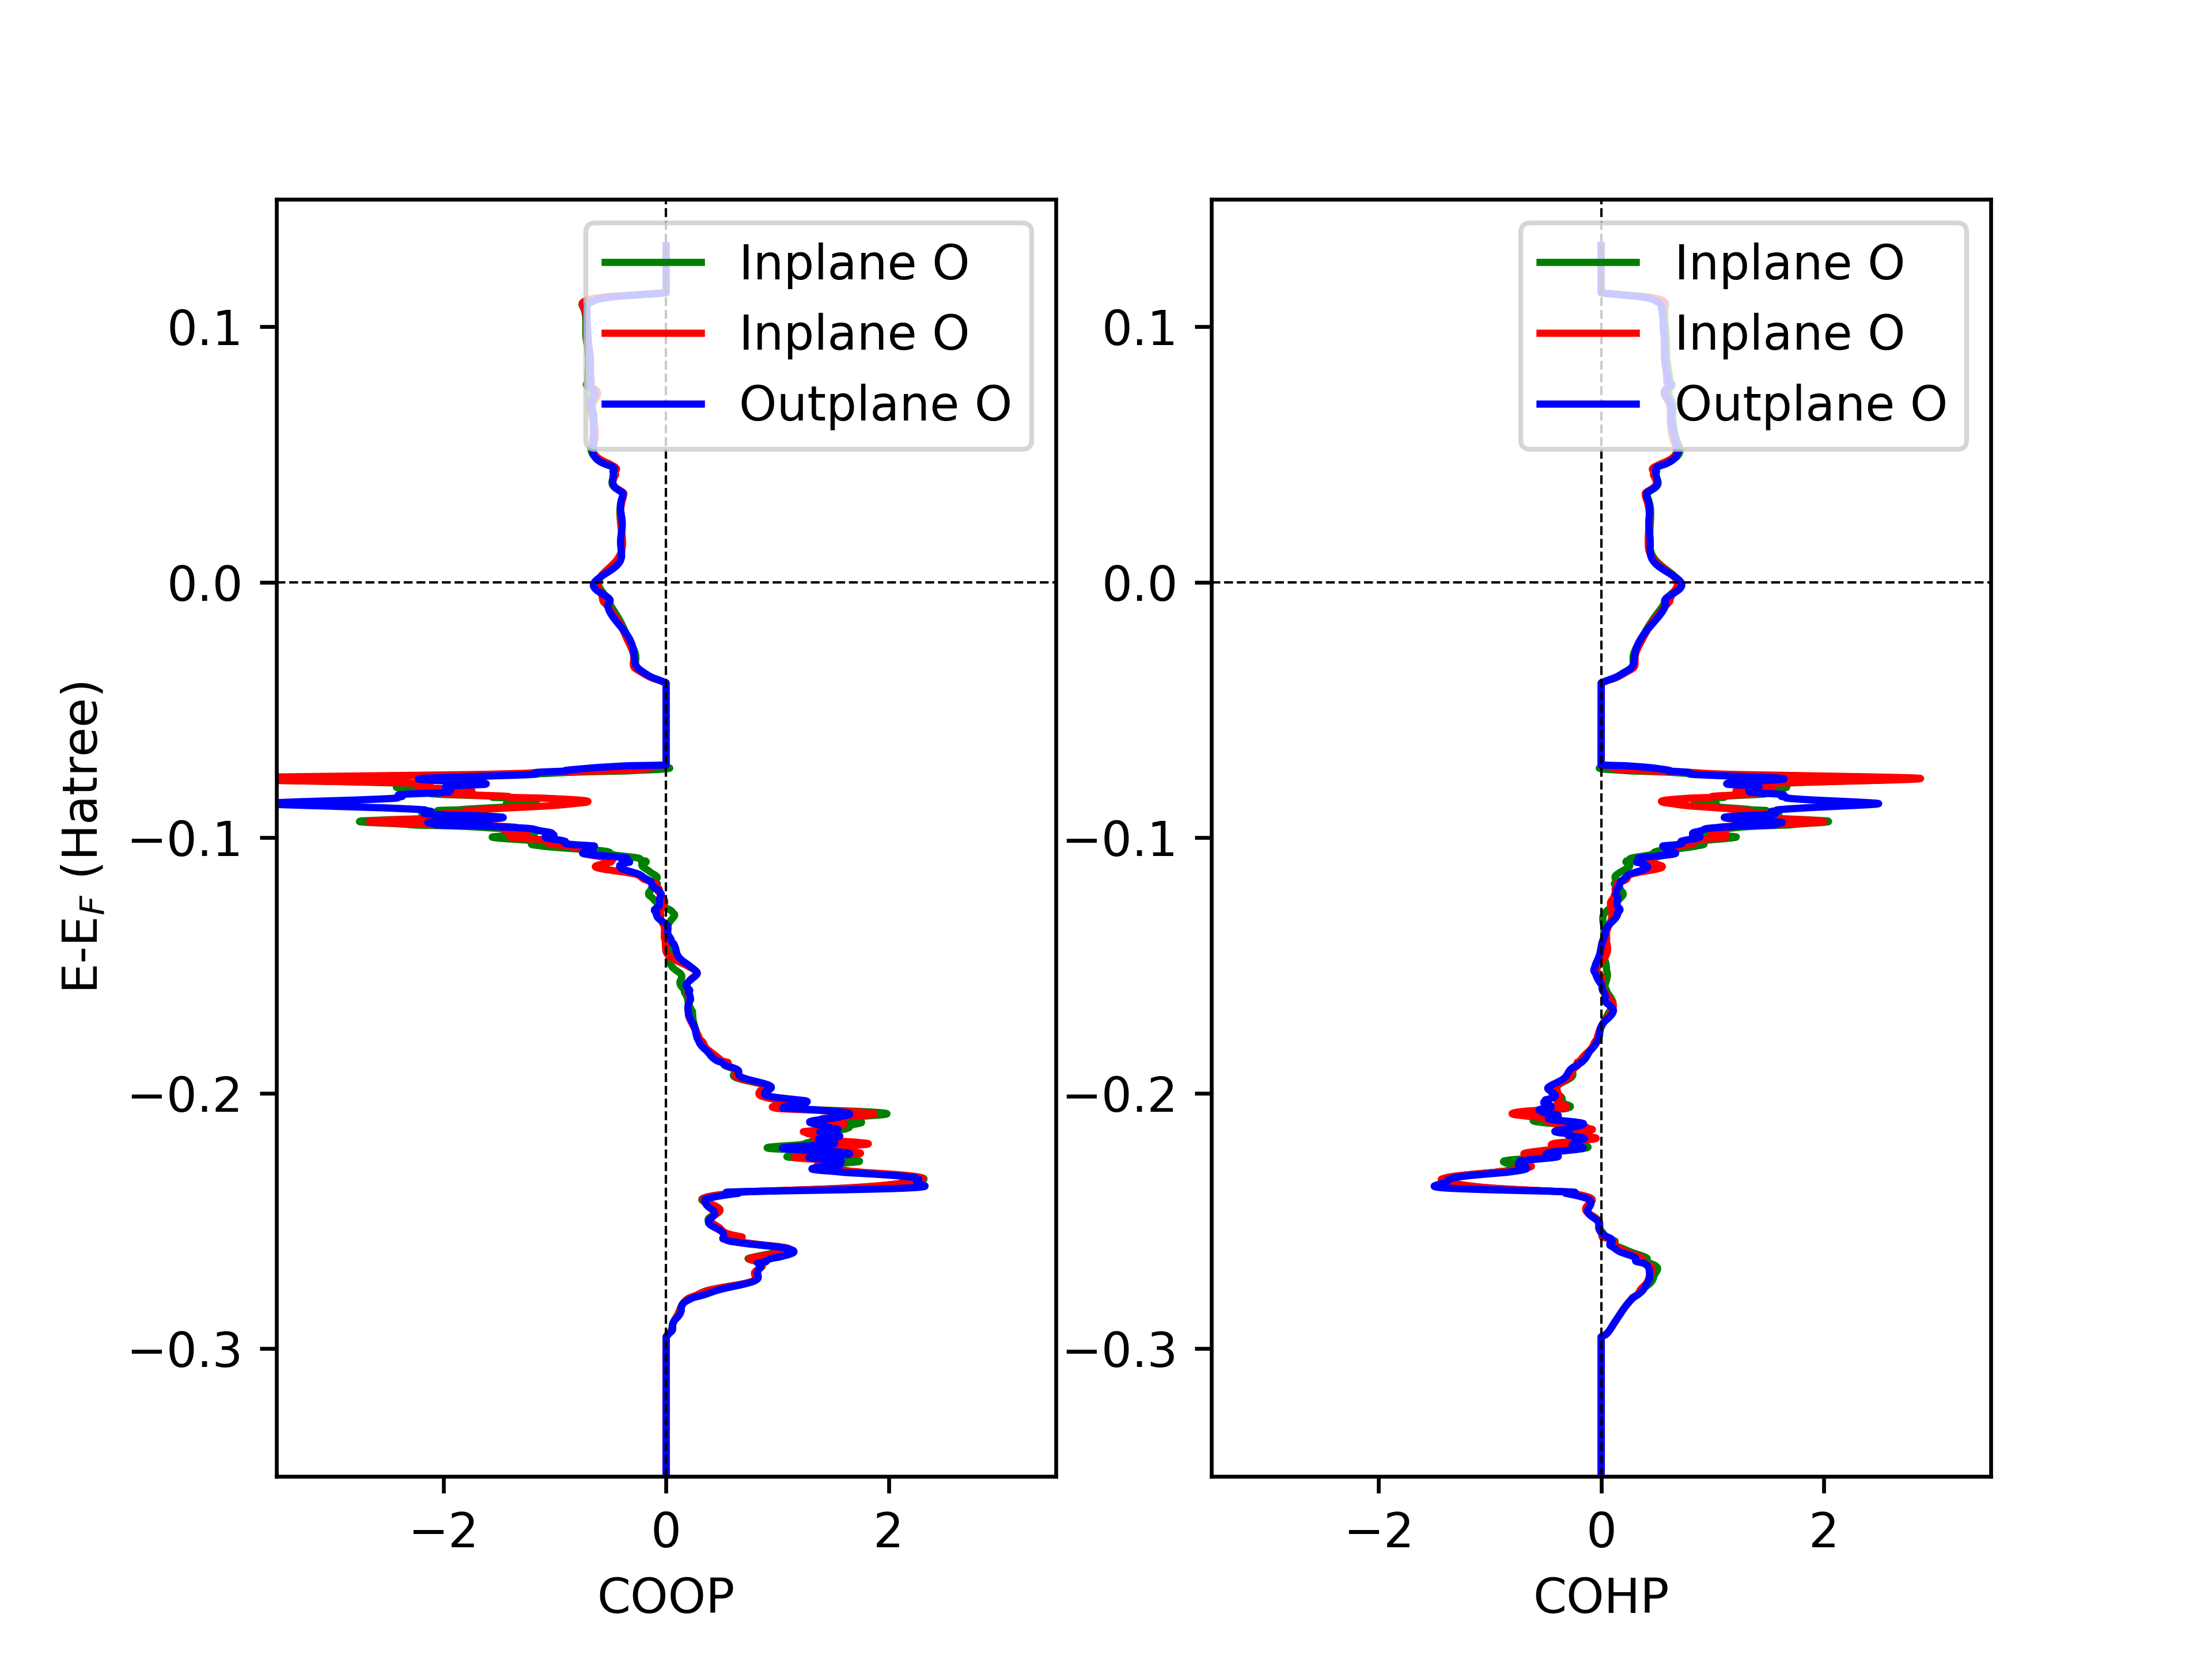
    Hi Prof. Erba, Thanks for the reply. From the file, I could see there are value from energy " -7.9324E-01" (Line No: 81826). I could plot the data using Python, Xmgrace. With CRYSPLOT somehow the data loading is slow and my browser hangs so I am not able to check it. But definitely there is data. I requested for COHP between Ni and three nearby oxygen. In the file there are 5 columns, so if I am not mistaken it corresponds to "Energy - Next three columns in the order I request - Final Column of DOS ?" (Because the last column data when plotted matches the DOS) Thanks & Regards, Rams
  • SCF fails spinlock with POB-DZVP-REV2

    Bug Reports
    10
    0 Votes
    10 Posts
    2k Views
    aerba
    Hi, @jquertin said in SCF fails spinlock with POB-DZVP-REV2: In the case of SPINLOCK, the manual clearly explain that the NSPIN value is the difference in number of alpha and beta electrons. For SPINLOC2, the text only refers to the spin while the table gives the same definition for SPIN as for NSPIN (in SPINLOCK). The argument SPIN of SPINLOC2 still represents a number of electrons, as in SPINLOCK. @jquertin said in SCF fails spinlock with POB-DZVP-REV2: Furthermore, in the calculation, if using SPINLOC2 with 6 or 6.0 as the spin (as defined in the table), crystal defaults to SPINLOCK. That's right. SPINLOC2 requires a non integer argument. For integer arguments it reduces to SPINLOCK. @jquertin said in SCF fails spinlock with POB-DZVP-REV2: In short, if I define SPINLOC2 SPIN as 3 (1/2 * 6) or 3.0, crystal defaults to SPINLOCK with NSPIN 3 which is actually half of what I want. In both SPINLOCK and SPINLOC2, the argument is meant as a number of electrons. Thus, if you have 6 extra up electrons with respect to down electrons, the input value should be 6, not 3. For integer values, SPINLOC2 is of no use. Hope this clarifies things a little,
  • ECHG in XY, XZ and XY-Z plots

    Electron Charge Density Analysis
    3
    0 Votes
    3 Posts
    558 Views
    Drmajouri2025
    Thank you, Dr. Alessandro Erba, for the helpful suggestion regarding the use of the COORDINA option. Your example for defining the XY plane was clear and easy to follow — I’ll definitely apply this approach in my setup. Thanks again for your guidance! Cheers
  • 0 Votes
    7 Posts
    1k Views
    QMQDCHEM
    Everything works with mpiifx, thank you very much!
  • SMALLDIST keyword

    Single-Point Calculations
    3
    0 Votes
    3 Posts
    679 Views
    ioikonom
    Hi Giacomo, Many thanks for your answer and suggestions. It works and will do a double check for the geometry. Have a good day, Ilias
  • 0 Votes
    8 Posts
    2k Views
    GiacomoAmbrogio
    Hi Chris, The reason you only see a few k-points printed is indeed that the printing options are not fully supported in parallel execution. In a parallel run, each MPI process handles a subset of the k-points, and only process 0 writes to the output file. As a result, you’ll only see the k-points assigned to process 0, which can be just a few, or even none, depending on how they are distributed. By default, the code distributes k-points in a round-robin fashion starting from the last process, so the gamma point is typically assigned to the last process and therefore never printed in a parallel execution. This behavior applies to both options 66 and 67. Try running the following input: NEWK 4 4 1 2 66 999 67 999 END However, you’ll need to run it on a single process for the eigenvectors to be printed correctly. If the calculation is too expensive to run on a single process, we may need to find an alternative approach to extract those values (though that won’t be straightforward).
  • 0 Votes
    5 Posts
    867 Views
    esmuigors
    Dear prof. Erba, Have You got any time to look into the source maybe? Quite interestingly, I have recently noticed that in some other QC code values for the components change a lot (by up to 60%) from run to run, except for the dipole orientation (when the molecular dipole is aligned with z axis). Is (or could be) the reorientation You are referring to connected with this? Seems like something similar to the struggle fir gauge invariance in NMR calculations...
  • 0 Votes
    2 Posts
    501 Views
    GiacomoAmbrogio
    Hi, We provide a distribution of precompiled object files that can be linked on this architecture. You should be able to build and run CRYSTAL23 successfully using those. We do not have specific performance optimization data or configuration recommendations for the KNL platform. Performance is highly dependent on the system setup and workload characteristics, so we recommend running a few short benchmarks to identify the best configuration for your case. As regards OpenMP, the optimal number of threads depends on your memory limitations; however, we recommend not exceeding 8 threads per MPI process. Let me know if you have any further questions.
  • BILLY software

    Basis Sets
    3
    0 Votes
    3 Posts
    688 Views
    Igor12
    Thank you!